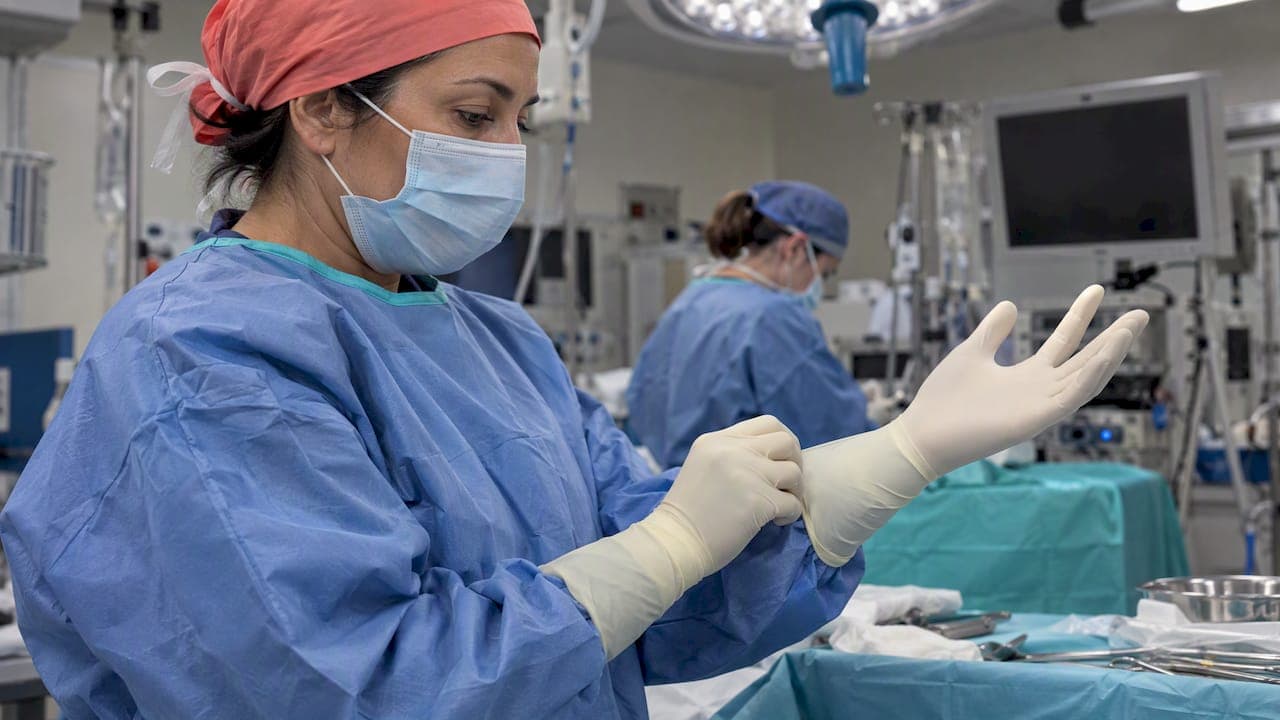
Double Gloving Guidelines: What U.S. Teams Should Follow
What CDC, OSHA, AORN and WHO say about double gloving, when to do it, how to don two pairs, and what procurement teams should stock for compliance.
Managing medical equipment import regulations is the process of ensuring medical devices comply with all legal, registration, and quality standards before and during importation to guarantee market access and patient safety. For regulatory officers and procurement managers, this process involves far more than clearing customs. It requires coordinating device classification, establishment registration, licensing, and documentation across multiple health authorities, including the FDA, the EU Medical Device Regulation framework, and Australia’s Therapeutic Goods Administration. Getting any one of these steps wrong delays shipments, exposes your organization to legal liability, and puts patient care at risk.
The first step in any compliant import is understanding your device’s classification. The FDA divides medical devices into Class I, II, and III based on risk level. Class I devices carry the lowest risk, while Class III devices require the most rigorous premarket approval. Getting the classification wrong changes your entire regulatory pathway, your duty rates, and your documentation requirements.
Before any shipment crosses a border, the following prerequisites must be in place:
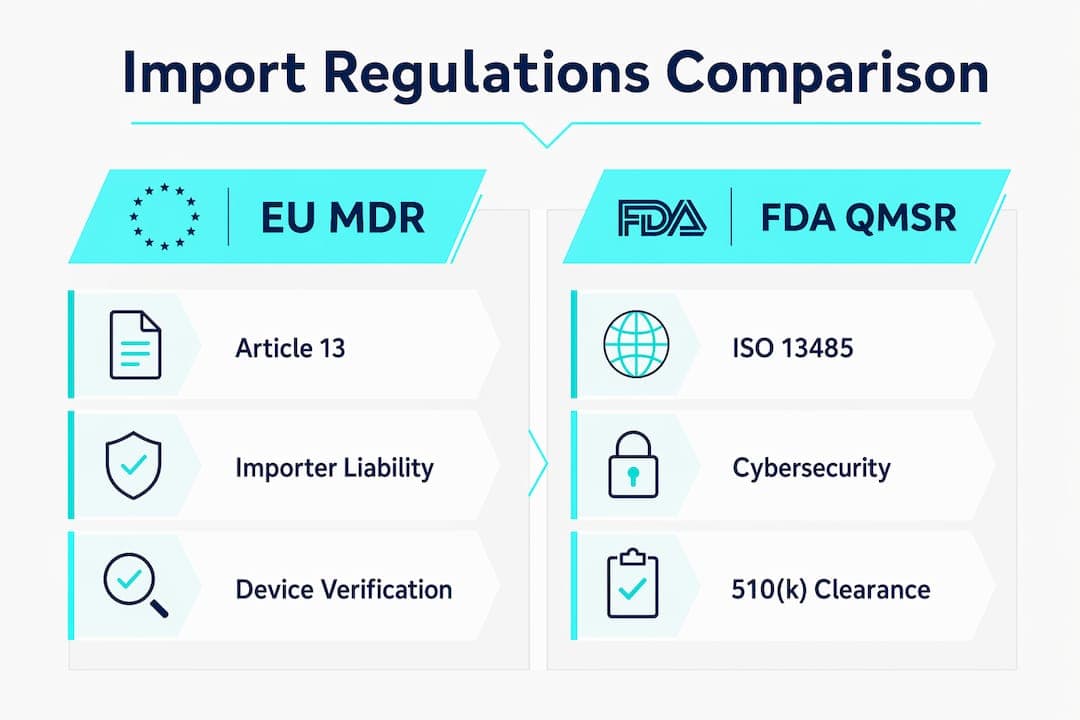
For EU imports, EU MDR Article 13 mandates that importers fulfill 10 specific obligations, including verifying device conformity, registering in EUDAMED as an Economic Operator, and maintaining complaint and non-conformity registers. This is not a paperwork formality. Importers under EU MDR carry joint and several liability for defective devices alongside manufacturers.
For Australia, the Therapeutic Goods Administration requires all devices to be listed on the Australian Register of Therapeutic Goods (ARTG) before importation. Skipping this step carries severe consequences. TGA penalties include up to 5 years imprisonment and fines of up to 5,000 penalty units, with each unit valued at $330 as of 2026.
Pro Tip: Map your device classification and registration requirements for each target market before finalizing procurement timelines. Registration timelines vary widely, and aligning them with your shipping schedule prevents costly holds at the border.
A compliant import follows a defined sequence. Skipping steps or running them in parallel without coordination is the most common source of delays.
Pro Tip: Build a registration calendar that tracks approval deadlines, renewal dates, and shipment windows for every device and every market. A missed renewal date can trigger an import hold just as damaging as a failed inspection.
The most frequent compliance failures are not technical. They are organizational. Procurement teams often treat medical device imports like standard freight, which means they miss the regulatory “second gate” that health authorities control after customs clearance.
Shipments arriving before device registration approval is complete are the leading cause of import holds. Customs clearance alone does not authorize a medical device for sale or use. Health authority approval is a separate and mandatory step that must precede the shipment, not follow it.
The following mistakes appear repeatedly across markets:
Non-compliance consequences extend beyond delays. Shipment detention, product destruction, import bans, and civil or criminal penalties are all possible outcomes depending on the jurisdiction and the severity of the violation.
Understanding the differences between major regulatory frameworks helps procurement managers build a global compliance strategy rather than reacting market by market.
| Market | Key Requirement | Importer Obligation | Notable Risk |
|---|---|---|---|
| United States (FDA) | 510(k) or PMA clearance; QMSR compliance from february 2026 | Designate U.S. Agent; annual registration | Cybersecurity documentation now mandatory |
| European Union (EU MDR) | CE marking; EUDAMED registration | Register as Economic Operator; 10 Article 13 obligations | Joint and several liability for defective devices |
| Australia (TGA) | ARTG listing before import | Sponsor registration; accurate labeling | Up to 5 years imprisonment and $1.65M in fines |
The FDA’s QMSR transition aligns U.S. requirements with ISO 13485:2016, which creates an opportunity. Organizations already certified to ISO 13485 have a compliance foundation that satisfies both FDA and EU MDR quality system requirements. Building your QMS on ISO 13485 reduces duplication across markets.
EU MDR Article 13 obligations place direct legal responsibility on importers, not just manufacturers. This shift means your procurement team must actively verify device conformity before accepting a shipment, not simply pass it through to the end user.
In the EU, combining the Authorized Representative and Importer of Record roles under a single entity is best practice. It eliminates coordination gaps between two parties that would otherwise need to exchange compliance documentation under time pressure.
Pro Tip: Use ISO 13485 certification as your global compliance baseline. It satisfies FDA QMSR, EU MDR quality system requirements, and TGA expectations simultaneously, reducing the cost of multi-market compliance.
Regulatory officers who manage imports across multiple markets need systems, not just checklists. The following approaches reduce errors and keep shipments moving.
Compliant medical device importation requires confirmed device classification, completed registration, valid importer licensing, and documented quality management before any shipment departs.
| Point | Details |
|---|---|
| Classify before you ship | Confirm device class and regulatory pathway for each market before procurement begins. |
| Registration must precede shipment | Health authority approval is a separate gate from customs clearance and must be completed first. |
| QMSR demands cybersecurity evidence | FDA inspections from 2026 onward require SPDF and SBOM documentation embedded in the QMS. |
| EU MDR creates direct importer liability | Importers carry joint legal responsibility for defective devices under Article 13 and Article 10(16). |
| ISO 13485 is your global baseline | Certification satisfies FDA, EU MDR, and TGA quality system requirements simultaneously. |
The biggest gap I see in medical device import operations is not a documentation gap. It is a mindset gap. Procurement teams are trained to move goods efficiently. Regulatory teams are trained to read frameworks. Neither group, on its own, is trained to do both at the same time under deadline pressure.
The 2026 QMSR transition is a concrete example of this problem. The cybersecurity requirements under Section 524B are not new in concept, but they are new as a hard enforcement priority. I have watched organizations spend months preparing their 510(k) submissions without once asking whether their software-enabled device had a documented SBOM. That oversight does not show up until an FDA inspector asks for it at the border.
The other lesson I keep returning to is the value of a single point of accountability. When the Authorized Representative and the Importer of Record are two different organizations, compliance gaps fall into the space between them. Every email thread, every handoff, every “I thought you handled that” is a potential delay. Consolidating those roles under one entity with healthcare expertise is not a luxury. It is the most practical risk reduction move available to a procurement manager working across multiple markets.
My honest advice: treat your import compliance program as a supply chain function, not a legal formality. Build it into your procurement calendar, fund it with real resources, and review it every time a regulation changes. The organizations that do this do not scramble when the FDA updates its guidance. They adjust and move on.
— QB
Healthcare procurement managers need suppliers who understand regulatory standards, not just product catalogs. Queenssurgical serves clinics, hospitals, and healthcare facilities across the Americas with a medical supply catalog built around quality and compliance.
Whether you are sourcing protective equipment like the CPE Thumb Loop Isolation Gown or the Full Length Face Shield, Queenssurgical stocks products that meet the standards your facility requires. Every product in the catalog is selected with healthcare-grade quality in mind, giving procurement teams a reliable source for compliant consumables and protective supplies. Visit Queenssurgical to browse the full catalog and connect with a team that understands what healthcare procurement actually demands.
Medical device import regulations are the legal requirements set by health authorities, such as the FDA, EU MDR, and TGA, that govern device classification, registration, licensing, and documentation before a device can legally enter a country.
Required documents typically include a commercial invoice, packing list, bill of lading, certificate of conformity, premarket clearance letter, and, for software-enabled devices, cybersecurity documentation such as an SBOM.
The most common cause of import holds is a shipment arriving before device registration is approved or the importer lacking the required medical device import license. Customs clearance does not substitute for health authority approval.
The QMSR transition, effective february 2, 2026, aligns FDA requirements with ISO 13485:2016 and adds mandatory cybersecurity documentation, including SPDF and SBOM, as part of quality management system compliance for all regulated devices.
The importer of record is the legally responsible party for ensuring a medical device meets all regulatory requirements at the point of entry. In the EU, this role now carries direct legal liability for device conformity under EU MDR Article 13.
What CDC, OSHA, AORN and WHO say about double gloving, when to do it, how to don two pairs, and what procurement teams should stock for compliance.
Mayo and Metzenbaum scissors look alike but do different jobs. See how blades, shanks and tips differ, when to use each, and how to care for them.
Sterile water or saline? Learn the difference in tonicity, safe uses, wound irrigation evidence, and how to choose irrigation and dressing supplies.